Research
Our research spans several interdisciplinary projects in computational systems biology. We use nonlinear dynamics, biophysics, data science, and bioinformatics to understand the behavior of biochemical networks and the principles of multicellular self-organization, and we work closely with experimental labs to test our models and theories.
Self-organization Stress response Differentiation Information-processing trade-offs Viral cell-fate decisions PK/PD modeling
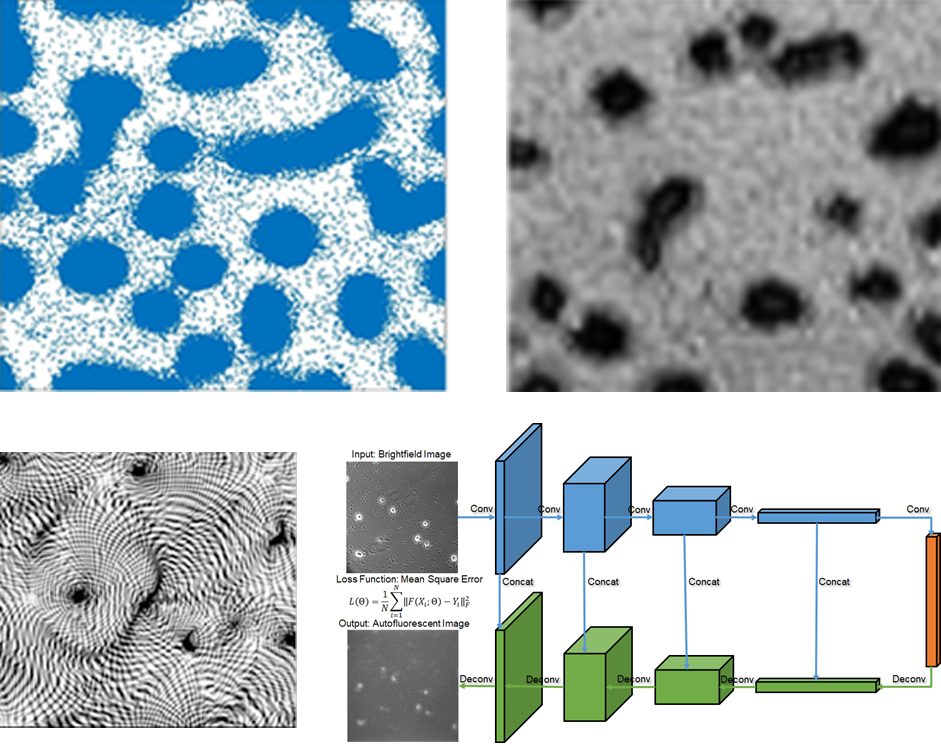
Mechanisms of microbial self-organization
How cells self-organize into complex tissues is one of the great puzzles of developmental biology, and a hallmark example of emergent behavior: complex patterns arising from simpler interacting components. Even the best-studied model systems still lack an understanding of the emergent properties that bridge developmental phenomena from molecules to cells, tissues, and whole organisms.
Biofilms formed by the soil bacterium Myxococcus xanthus are an excellent model system for emergent behavior. Under starvation, an M. xanthus biofilm starts a developmental program in which cells aggregate into mounds and then differentiate into distinct cell types. Complementing experimental research, we use computational biophysics and engineering approaches to build mathematical models of these phenomena, and we use recent advances in machine learning to connect the collective behavior of cells in a biofilm to their underlying genetic networks.
Selected papers
- Zhang J, et al. Deep learning framework for quantifying self-organization in Myxococcus xanthus. PNAS (2026)
- Murphy P, et al. Cell behaviors underlying Myxococcus xanthus aggregate dispersal. mSystems (2023)
- Cotter CR, et al. Data-driven modeling reveals cell behaviors controlling self-organization during Myxococcus xanthus development. PNAS (2017)
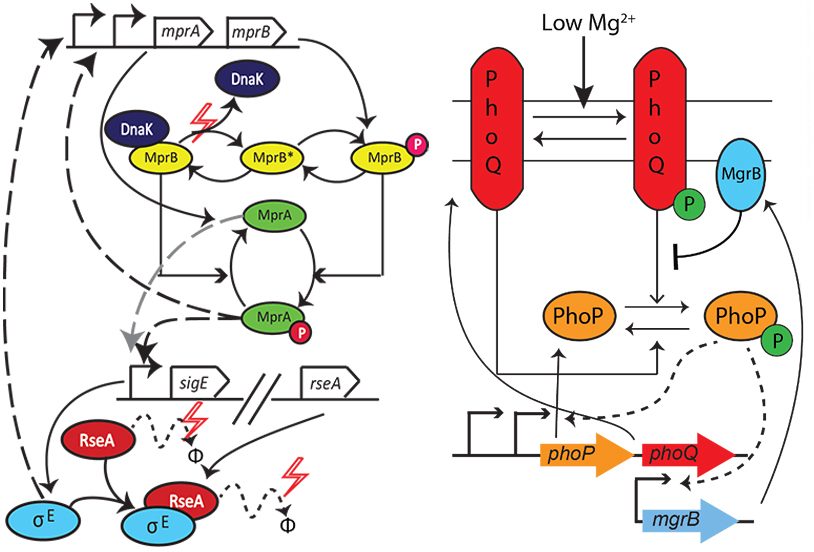
Evolutionary design principles of bacterial stress response
When exposed to harsh environments, antibiotics, or the immune response, bacteria survive by switching their gene-expression program into a “stress-response” mode. We want to understand how the biochemical networks that sense and respond to stress are organized in specific model systems, and to formulate general principles of stress response across bacterial species. Although their regulatory core is conserved, these networks vary in architecture. We look for the evolutionary design principles behind them by correlating these variations with different regulatory demands, focusing on bacterial sigma factors and two-component systems, two widespread mechanisms of master-level regulation of bacterial gene expression.
The stress-response network of Mycobacterium tuberculosis is critical for the survival of this intracellular pathogen within the host. To survive infection, M. tuberculosis must sense the stress conditions generated by the host immune system and adapt by reprogramming its gene expression and metabolism. With experimental microbiologists, we study how combinations of transcriptional and post-translational interactions shape the dynamics of stress-response networks.
Selected papers
- Rao SD, et al. Chaperone-mediated stress sensing in Mycobacterium tuberculosis enables fast activation and sustained response. mSystems (2021)
- Rao SD, Igoshin OA. Overlaid positive and negative feedback loops shape dynamical properties of PhoPQ two-component system. PLOS Comput Biol (2021)
- Narula J, Tiwari A, Igoshin OA. Role of autoregulation and relative synthesis of operon partners in alternative sigma factor networks. PLOS Comput Biol (2016)
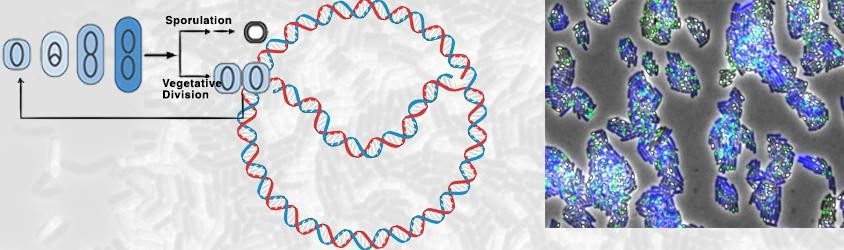
Functional requirements of cellular differentiation
Executing a differentiation program requires cells to weigh many internal and external cues and respond with the appropriate gene-expression program. The soil bacterium Bacillus subtilis has long been a model for prokaryotic differentiation. We use mathematical modeling to gain a systems-level understanding of how B. subtilis cells sense starvation and decide whether to start a unicellular (sporulation) or multicellular (biofilm) differentiation program.
We investigate how internal and external cues are dynamically encoded in the activity of master regulators of differentiation, and how different dynamics of a single master transcriptional regulator — that is, how its activity changes over time — can direct cells to alternative fates.
Selected papers
- Zhang Y, et al. Biophysical modeling reveals the transcriptional regulatory mechanism of Spo0A, the master regulator in starving Bacillus subtilis. mSystems (2025)
- Palma CSD, et al. Changes in Spo0A~P pulsing frequency control biofilm matrix deactivation. PLOS Comput Biol (2025)
- Narula J, et al. Slowdown of growth controls cellular differentiation. Mol Syst Biol (2016)
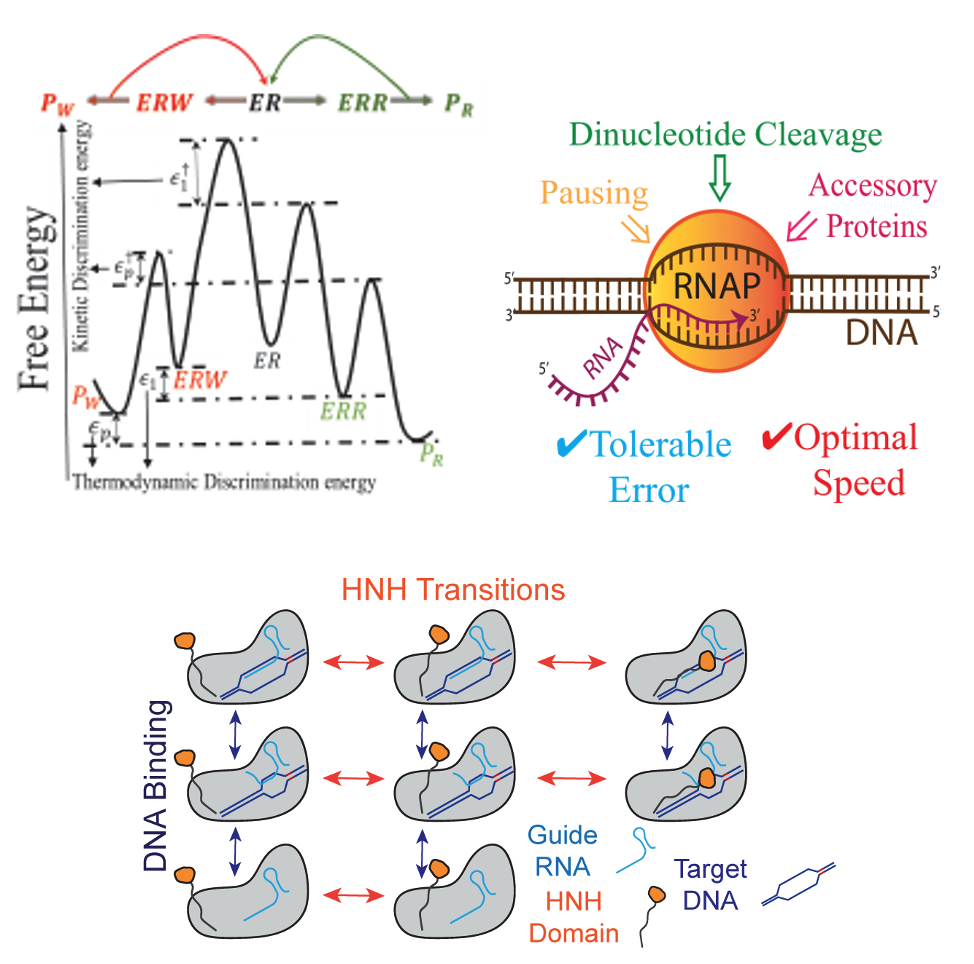
Trade-offs in non-equilibrium biochemical information processing
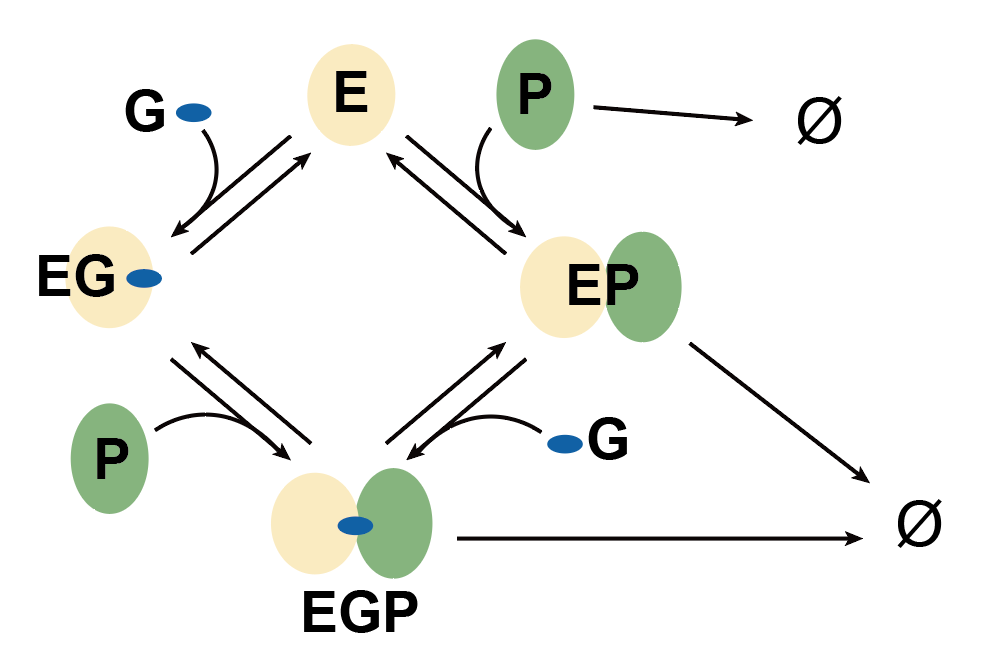
The ability of enzymes and biochemical networks to read and propagate molecular information is key to the complexity of life. Yet thermal fluctuations and fluctuations in molecular copy numbers place fundamental limits on the precision of information processing. We use kinetic and statistical-physics approaches to understand how enzymatic mechanisms and free-energy landscapes evolved to balance speed, accuracy, and energy dissipation.
We apply these methods to DNA replication, transcription, and translation, as well as to gene regulation and chromatin dynamics. Our work on transcriptional fidelity shows how sequence-dependent kinetics, proofreading, and pausing set transcriptional error and speed. For the nuclease CRISPR-Cas9, we built a kinetic model of substrate selection, determined its parameters from single-molecule and high-throughput data, and used residence-time distributions to match measured conformational dynamics. This revealed intrinsic trade-offs between on-target cleavage speed and off-target specificity that can inform Cas9 engineering. We also study how chemical inhibitors and modulators change enzyme specificity, and we use data-driven parameter inference to connect theory with experiments. Together, these studies reveal fundamental constraints on biochemical information processing, with implications for synthetic biology and disease modeling.
Selected papers
- Midha T, Kolomeisky AB, Igoshin OA. Kinetic mechanisms for the sequence dependence of transcriptional errors. PNAS (2025)
- Hecht AD, Igoshin OA. Kinetic mechanism for fidelity of CRISPR-Cas9 variants. J Phys Chem Lett (2025)
- Banerjee K, Kolomeisky AB, Igoshin OA. Elucidating interplay of speed and accuracy in biological error correction. PNAS (2017)
Mathematical modeling of pharmacokinetics and pharmacodynamics
Rational design of drug delivery requires precise knowledge of how a drug distributes through the body (pharmacokinetics, PK) and how it affects its target tissue (pharmacodynamics, PD). For protein therapeutics such as monoclonal antibodies and recombinant cytokines, these relationships are further complicated because a drug’s PD can affect its PK. And while preclinical animal models provide crucial insights, predicting how PK/PD data translate to humans remains a major challenge.
To address these complexities and improve predictive accuracy, we develop mathematical models built on data collected by our collaborators. We apply these models to understand and predict human treatment outcomes in promising areas such as immunotherapy, molecular glues, and GLP-1 receptor agonists.
Selected papers
- DeBonis J, Veiseh O, Igoshin OA. Uncovering the interleukin-12 pharmacokinetic desensitization mechanism and its consequences with mathematical modeling. CPT Pharmacometrics Syst Pharmacol (2025)
- Chae SJ, et al. Biophysical modeling elucidates mechanistic principles for rational molecular glue design. J Chem Inf Model (2026)
- Fell C, et al. Once yearly cell-based therapy for sustained and dose-tunable delivery of monoclonal antibodies. Science Advances (2026)